Protein Molecular Dynamics Simulation Tutorial
Published on October 11, 2026
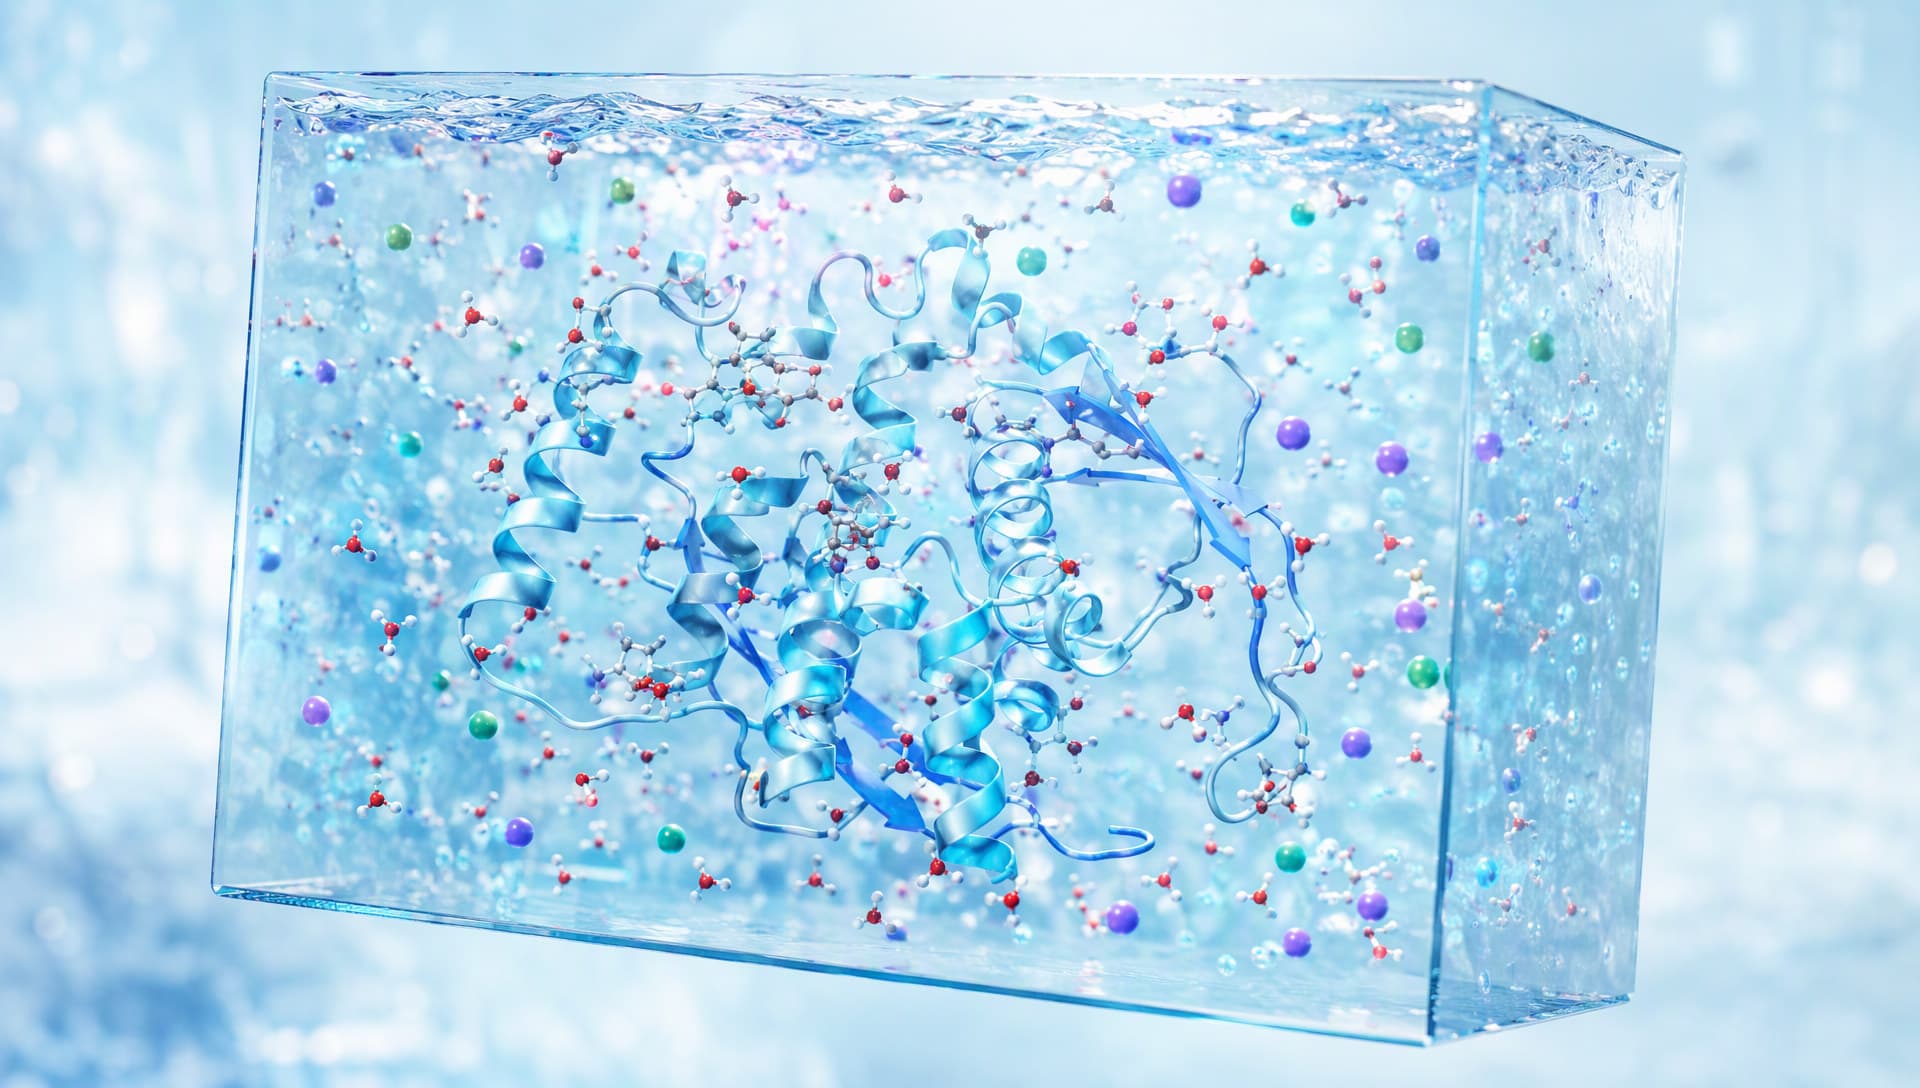
The solvated simulation box reveals how protein conformations evolve over time
Category: Computational Biology | Structural Biology | Protein Engineering
A crystal or predicted structure is a valuable starting point, but it does not show how a protein continuously explores conformations in solution. An active-site loop may open and close, a mutation may reshape local flexibility, and a ligand may reorganize within its pocket. The practical goal is therefore not to produce an attractive movie. It is to build a traceable chain of dynamic evidence that answers a defined question. A useful protein molecular dynamics simulation tutorial must connect input quality, physical assumptions, sampling, and interpretation so that successful software execution is not mistaken for a scientific conclusion.
A Protein Molecular Dynamics Simulation Tutorial Should Begin with the Question
Simulation design changes with the decision at hand. A comparison of wild type and mutant proteins requires consistent structural models, matched parameters, independent replicates, and predefined local readouts. Testing whether a ligand pose persists adds ligand parameterization, protonation choices, pocket waters, and interaction occupancy. Investigating a flexible loop may require longer or enhanced sampling and state-based analysis. Before opening GROMACS, define the comparison, primary observables, acceptance criteria, and the experiment that would test the computational hypothesis.
Structure preparation is the first quality gate. Inspect missing residues, alternative conformations, chain identifiers, nonstandard residues, metals, cofactors, and crystallographic waters. Assign protonation states in the context of the intended pH and local environment. Long missing regions should not be rebuilt without a defensible model, while every water molecule should not be deleted automatically; a conserved pocket water can mediate a critical interaction. When only a raw sequence is available, protein identity and structural provenance should be established first. MatwingsVenus™(晓鹜™) can connect database retrieval from resources such as PDB and UniProt with structure preparation in the same conversational task, helping teams document inputs and unresolved choices before expensive computation begins.
The force field and water model should be selected as a compatible combination and kept consistent across comparisons. Standard amino acids are generally covered, whereas ligands, modified residues, metal centers, and covalent linkages often require additional parameterization and manual review. The periodic box needs sufficient padding, and ions should both neutralize the system and represent the intended conditions. There is no universal parameter set for every protein. Reproducibility depends on recording software versions, force fields, water models, box geometry, ionic strength, parameter sources, and all departures from the baseline protocol.
Each Stage Needs a Gate Before the Next One Starts
Energy minimization removes severe clashes and unfavorable contacts; it does not prove that the protein has become more biologically correct. Equilibration commonly proceeds through NVT to approach the target temperature and NPT to stabilize pressure and density. Early position restraints can allow solvent and ions to adapt around the protein before restraints are gradually released. Entry into production should depend on quality indicators rather than on completion of a fixed number of steps. Potential energy, temperature, pressure, density, and selected structural observables should all be inspected over an appropriate window.

The workflow links system building, equilibration, production, and quality control
A compact task specification is more useful than a loose collection of commands:
Goal: compare local stability of a wild type and a mutant under matched conditions
Inputs: two reviewed structures, one protonation policy, and identical cofactors
Workflow: minimization -> NVT -> NPT -> multiple independent production replicas
Primary readouts: global RMSD, local RMSF, key hydrogen-bond occupancy, clustering
Decision rule: report a comparison only when replicas agree and equilibrium is justified
Outputs: parameter record, quality-control plots, processed trajectories, analysis scripts
How MatwingsVenus™(protein agent)Connects the GROMACS Workflow
This is the type of connected workflow that MatwingsVenus™(晓鹜™) is designed to support. Its GROMACS toolset covers ligand preparation, simulation, analysis, custom scripts, and format conversion, with a human confirmation gate before compute-intensive execution. Researchers can describe the system, parameters, and expected outputs in natural language, review the proposed task, and then authorize the run. This does not remove scientific judgment; it reduces avoidable context loss when files, commands, and analysis steps move between separate tools.
Production length has no universal value outside a defined research question. A short trajectory can validate the workflow or reveal gross instability, but it cannot establish that conformational space has been sampled adequately. Multiple replicas with independent random seeds often provide stronger evidence because agreement can be evaluated directly. Slow transitions may require longer simulations or an enhanced-sampling method appropriate to the question. More nanoseconds do not automatically produce better evidence when parameters, equilibration, quality control, or replication remain weak.
Trajectory Analysis Must Answer the Hypothesis, Not Merely Produce Plots
RMSD measures displacement relative to a reference, but a plateau does not prove that the biologically relevant state is correct. RMSF describes residue-level fluctuations, yet its interpretation depends on the alignment and on domain motion. Radius of gyration and solvent-accessible surface area can reveal changes in compactness or exposure, but neither alone demonstrates improved stability. Hydrogen bonds, salt bridges, contact frequencies, and key distances are better suited to specific interactions. Clustering, principal component analysis, and free-energy landscapes can help identify dominant states and transitions. The right analysis combines a small set of complementary observables chosen before the trajectory is inspected.

Conformational ensembles and energy landscapes turn trajectories into testable insight
Trajectory processing also matters. Molecules crossing periodic boundaries must be made whole, the system should be centered, and the fitting group must match the question. Fitting an entire protein before interpreting a mobile domain can hide or exaggerate local motion. Showing only the smoothest replica is equally misleading. A reproducible protein molecular dynamics simulation tutorial should preserve raw trajectories, processing commands, index groups, scripts, software versions, and the exact segment excluded as equilibration, together with the reason for exclusion.
In protein engineering, simulation is best treated as evidence for prioritization rather than as a substitute for experiment. MatwingsVenus™(晓鹜™) can organize trajectory analysis alongside upstream database retrieval, functional-site identification, and downstream mutation assessment. A team can first determine whether a proposed mutation overlaps a catalytic or conserved region, then compare local interaction networks and state populations, and finally produce a shortlist for experimental testing. Computational outputs should remain labeled as predictions, while thermal stability, activity, affinity, or expression assays provide the endpoint that matches the engineering objective.
FAQ: Questions That Affect Interpretation
Does an RMSD plateau prove convergence? No. Examine stationarity of the observables tied to the hypothesis, compare distributions across time windows, test agreement among independent replicas, and ask whether relevant state transitions were sampled.
Must NVT and NPT always run for fixed durations? The common sequence is NVT for temperature stabilization followed by NPT for pressure and density, but duration should be governed by the system response and the intended production ensemble. Membrane proteins, metal centers, and large complexes may require more careful coupling and longer equilibration.
Which metrics belong in a protein molecular dynamics simulation tutorial? Work backward from the hypothesis. Global stability may combine RMSD, radius of gyration, and clustering; local flexibility may use neighborhood RMSF and secondary-structure changes; binding questions may prioritize contacts, hydrogen bonds, key distances, and, when justified, free-energy estimates with explicit method limitations.
Can an AI agent replace an experienced simulation scientist? Its more defensible role is to support planning, tool orchestration, input-output organization, and continuity across a research workflow. Researchers still need to approve the system and cost, diagnose anomalies, assess sampling, and connect computational observations to experimental evidence.
Conclusion
A reliable protein molecular dynamics simulation tutorial is not a command list. It is a reproducible process built around question definition, input review, transparent parameters, stage gates, replicated sampling, and hypothesis-led analysis. MatwingsVenus™(晓鹜™) helps connect database retrieval, structure preparation, GROMACS simulation, trajectory analysis, custom scripts, and subsequent protein engineering within one conversational workflow while preserving human approval and experimental-validation boundaries. The result is not merely a trajectory file, but an evidence chain that can guide the next design and validation cycle.